
CART
No products in the cart.
The 21 CFR Part 11 Toolkit gives you 105 editable templates for computer system validation and electronic records: 68 Word documents and 37 Excel workbooks. It covers 21 CFR Part 11, EU GMP Annex 11 (2011, in force) and every numbered clause of the July 2025 Annex 11 draft, with the validation master plan, URS, IQ, OQ and PQ protocols, traceability matrix, audit trail review and e-signature controls. Independent of FDA and the European Commission.
$99.00
✓ In stock — instant download after checkout
At a glance: the 21 CFR Part 11 Toolkit is an editable Microsoft Word and Excel documentation set for 21 CFR Part 11 and EU GMP Annex 11 (2011 revision, in force), arranged on the July 2025 Annex 11 draft. It covers the validation master plan and system qualification, GxP impact and risk assessment, supplier and SaaS assessment, data integrity (ALCOA+), audit trail review, electronic signatures and access control, and periodic review, backup and archiving. One-time price $99, instant download, licensed for internal use by one organisation. Implementing for several clients? The Consultant Package covers unlimited client engagements.
21 CFR Part 11 sets the conditions under which FDA accepts electronic records and electronic signatures in place of paper and ink. It applies whenever a record that an FDA regulation requires is kept, signed or submitted electronically. In the EU, GMP Annex 11 sets the matching expectations for every computerized system used in medicines manufacturing. Inspectors on both sides ask the same questions: is the system validated, who can change the data, is every change in the audit trail, and does a signature really belong to the person who applied it.
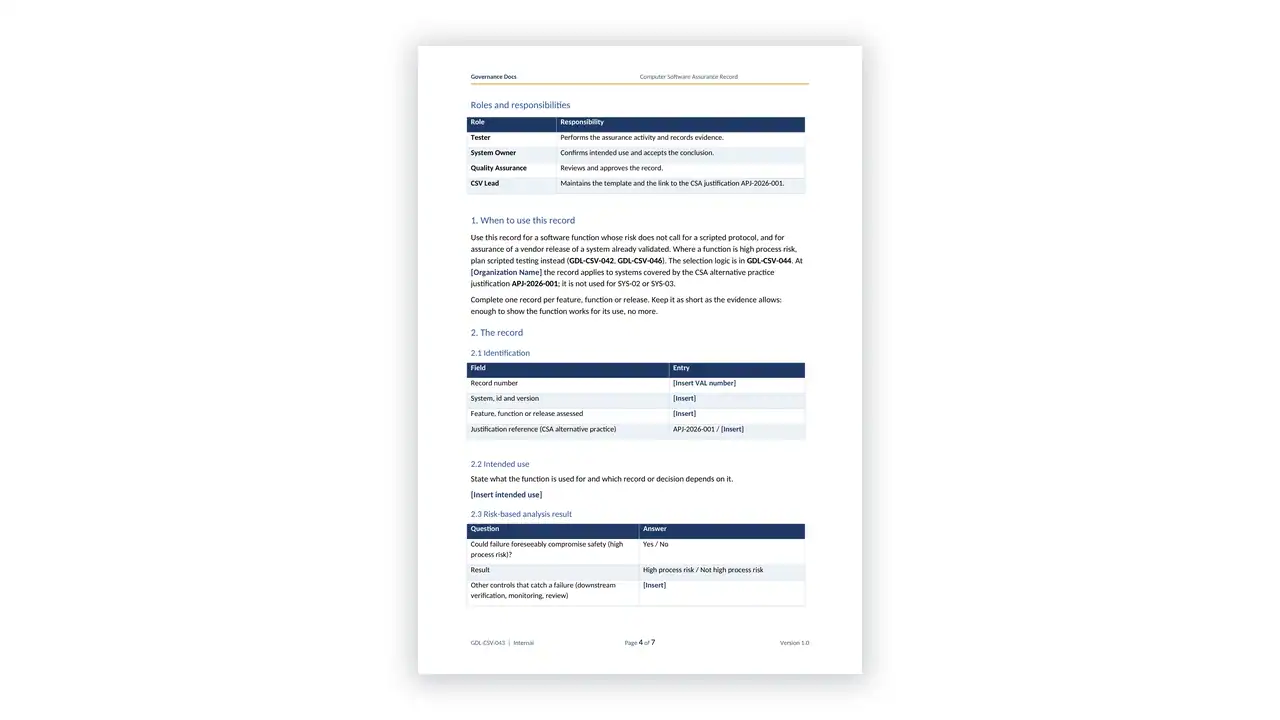
The 21 CFR Part 11 Toolkit (the Computer System Validation Toolkit) gives you the documents to answer those questions: the procedures that run the program, the validation deliverables for each system, and the registers and checklists that prove it to an inspector. Folders follow the section order of the July 2025 Annex 11 draft, with each Part 11 and 2011 Annex 11 requirement placed where its topic lives, so one structure serves US and EU inspections.
| Source | Status | What the pack covers |
|---|---|---|
| 21 CFR Part 11 | In force (US) | Every section from 11.1 to 11.300, broken into 38 requirement rows |
| EU GMP Annex 11, January 2011 revision | In force (EU) | The principle and all 32 numbered clauses |
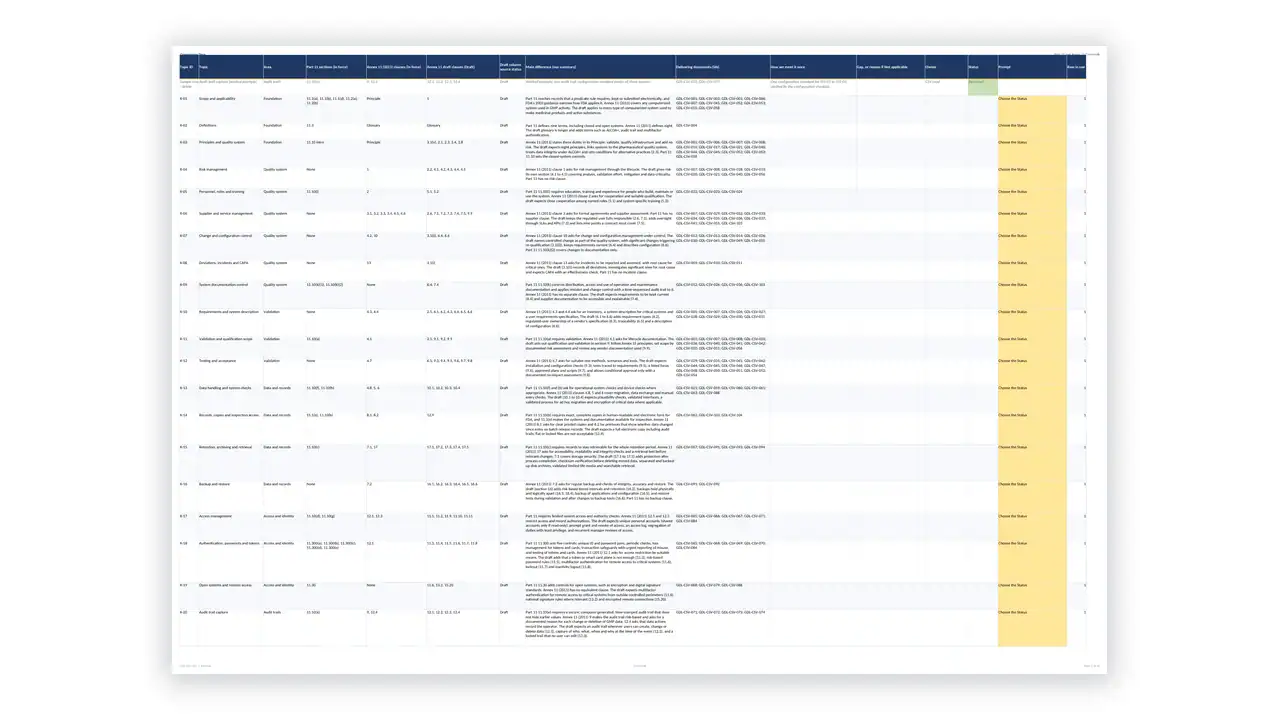
| Annex 11 consultation draft, July 2025 | Draft, not adopted | All 17 sections and all 111 numbered sub-clauses |
| Annex 22 (artificial intelligence) draft, July 2025 | Draft, not adopted | All 10 sections and 33 numbered sub-clauses |
| FDA Computer Software Assurance guidance, 3 February 2026 | Final guidance, device scope | Used for the risk-based assurance method |
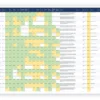
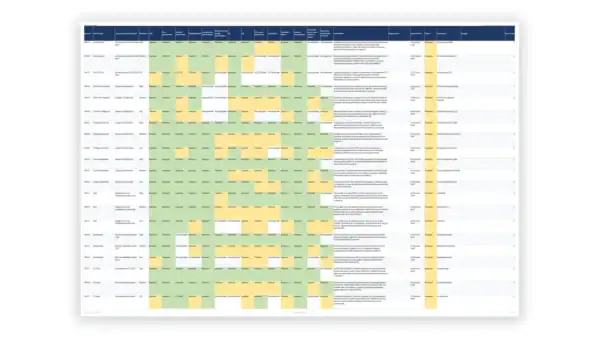
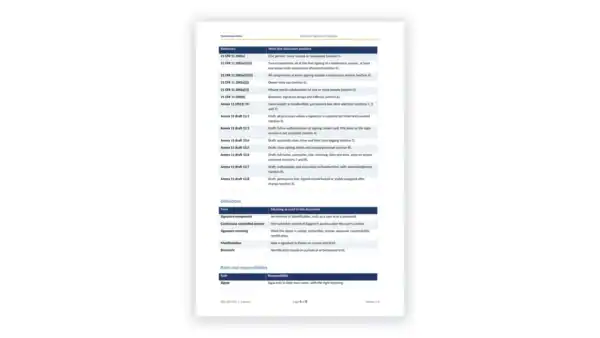
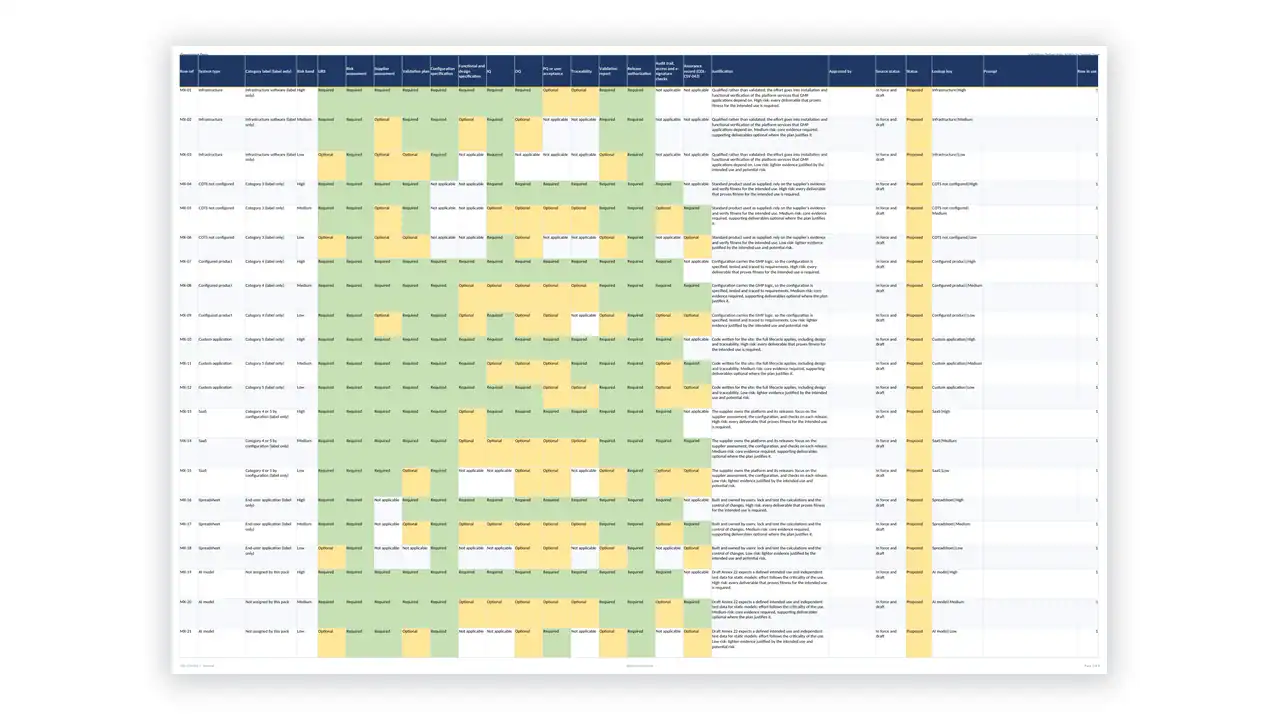
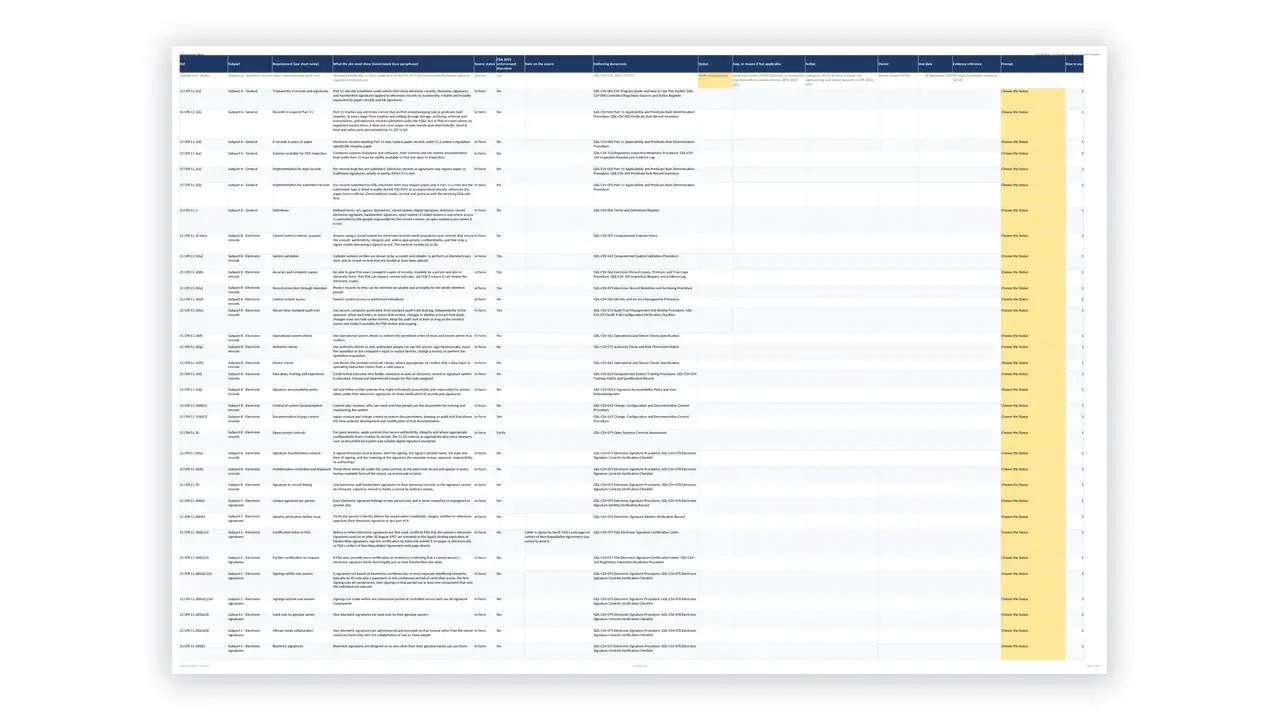
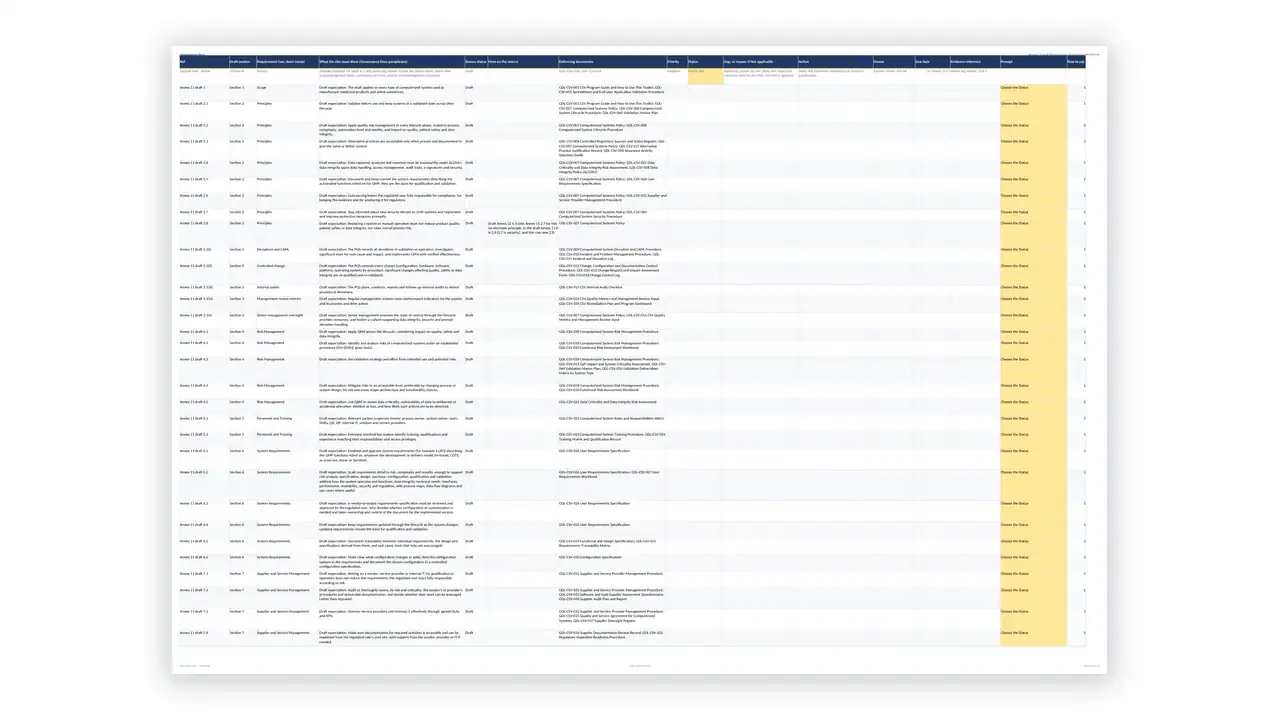
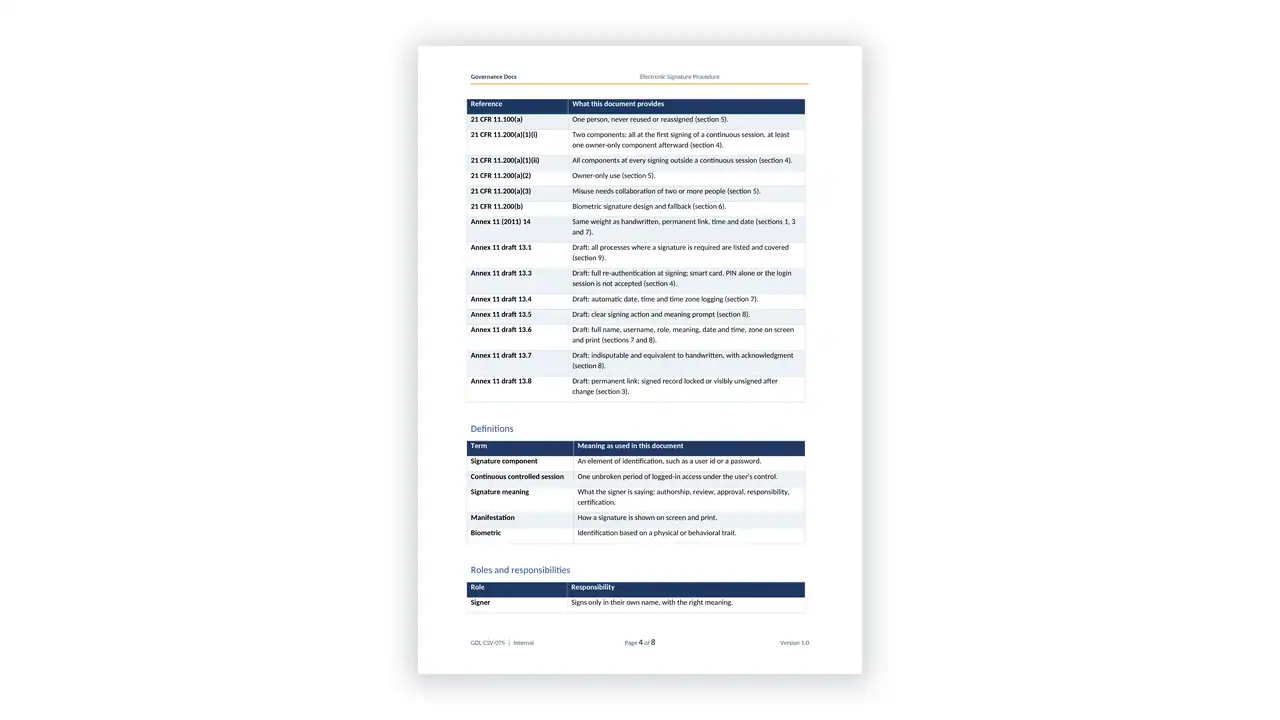
Every Word document opens with a table of the requirements it answers and where in the document each one is met. The build that produced this pack fails if any requirement of Part 11, the 2011 Annex 11 or either draft is left without a working document. That is a check we run, not a claim we make. The Requirement to Document Master Matrix lists all 233 requirement rows and the documents that deliver each.
The Commission consulted on a rewritten Annex 11, a new Annex 22 on AI and a revised Chapter 4 from 7 July to 7 October
drafts an adoption date, because none has been published. Instead:
the gap before the final text lands.
Production and Quality Management System Software”, replacing the version of 24 September 2025. The pack follows the 2026 text.
borrow its risk-based logic only with a documented justification, and the pack includes the Alternative Practice Justification Record for exactly that.
System Regulation took effect on 2 February 2026. The pack never treats it as current.
The AI documents cite 2.8 and say why.
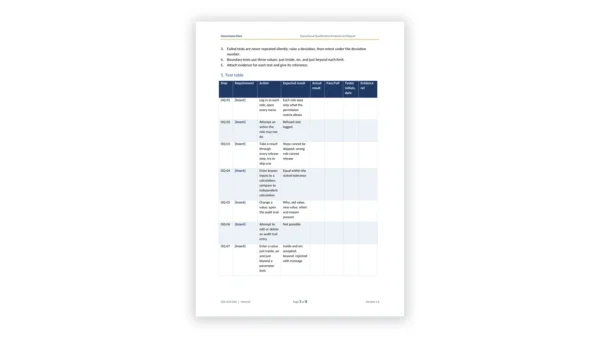
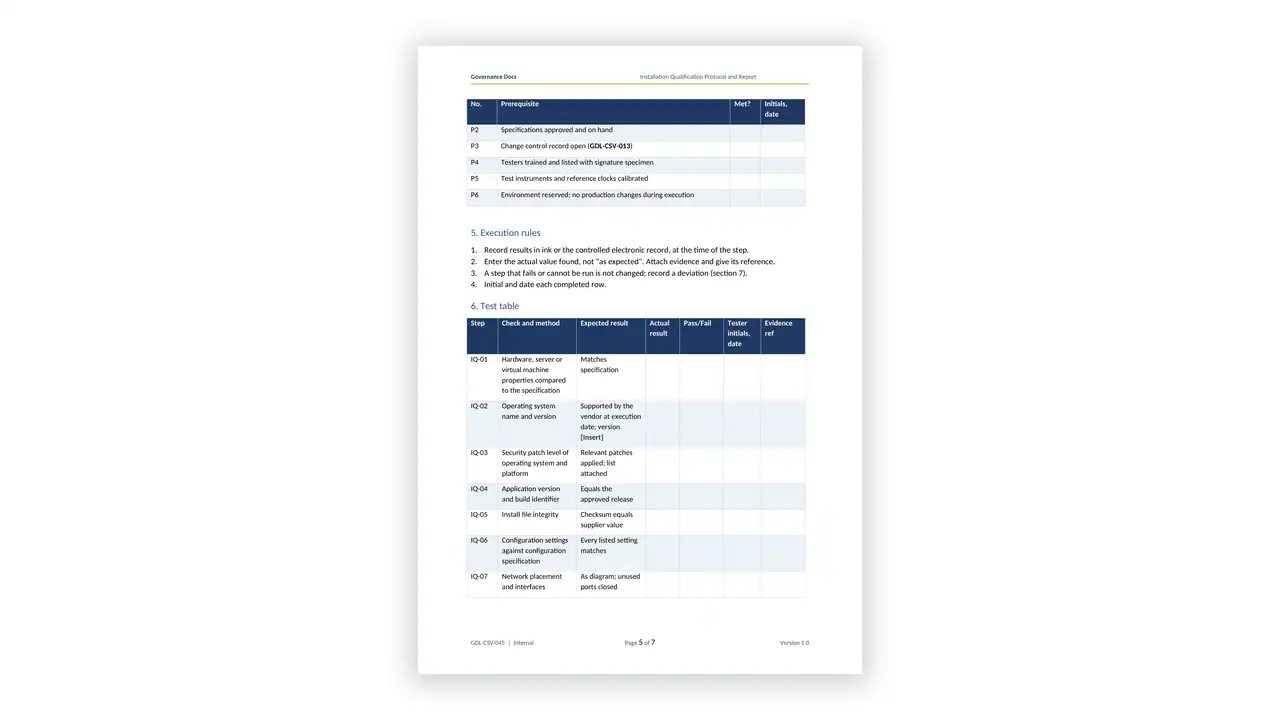
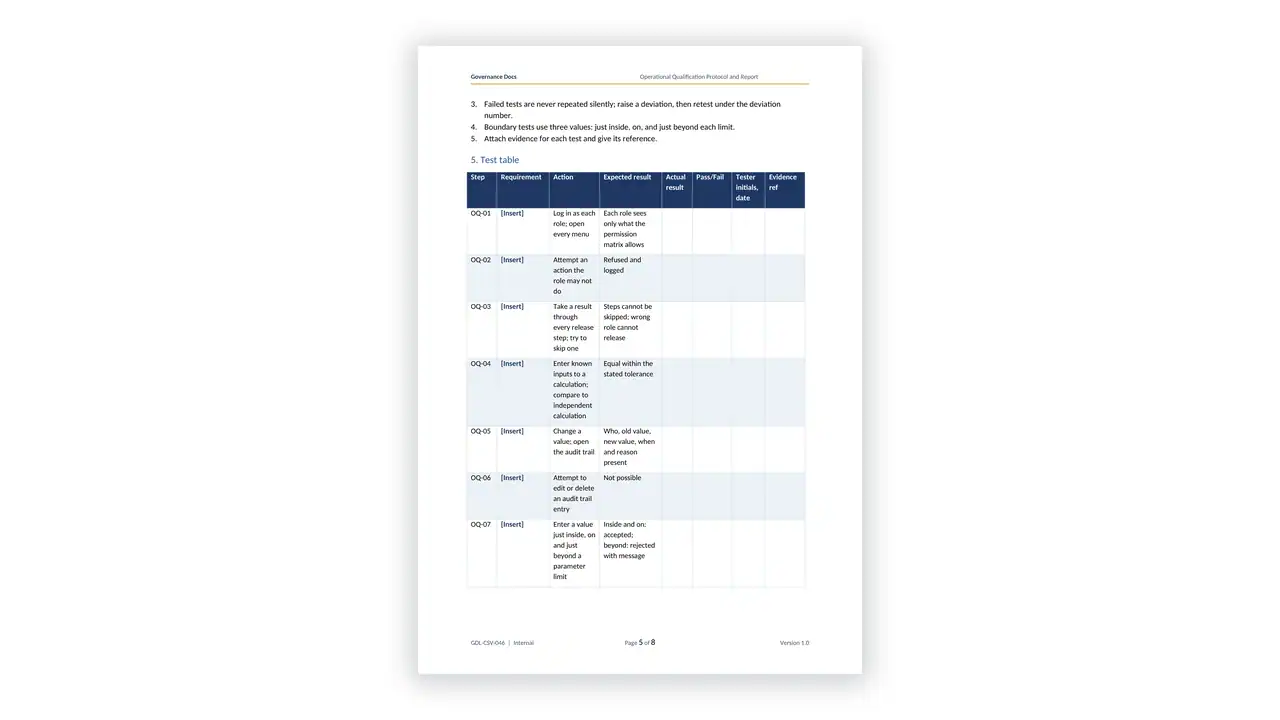
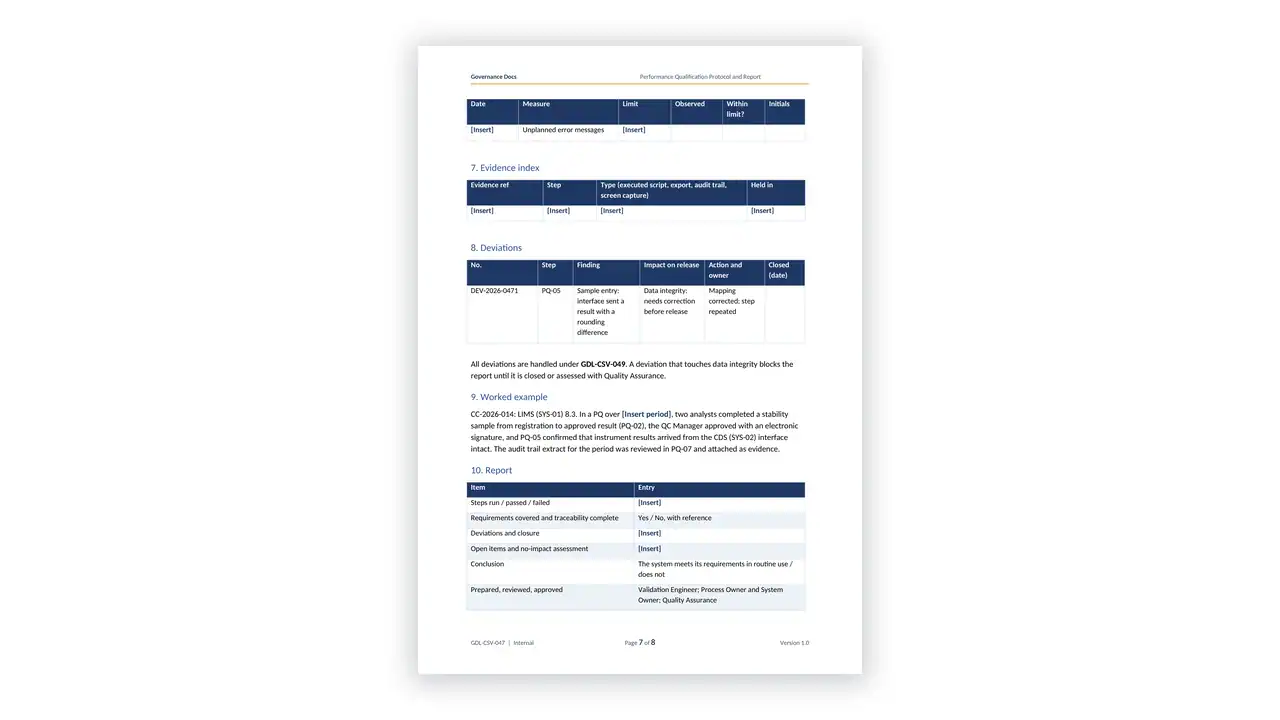
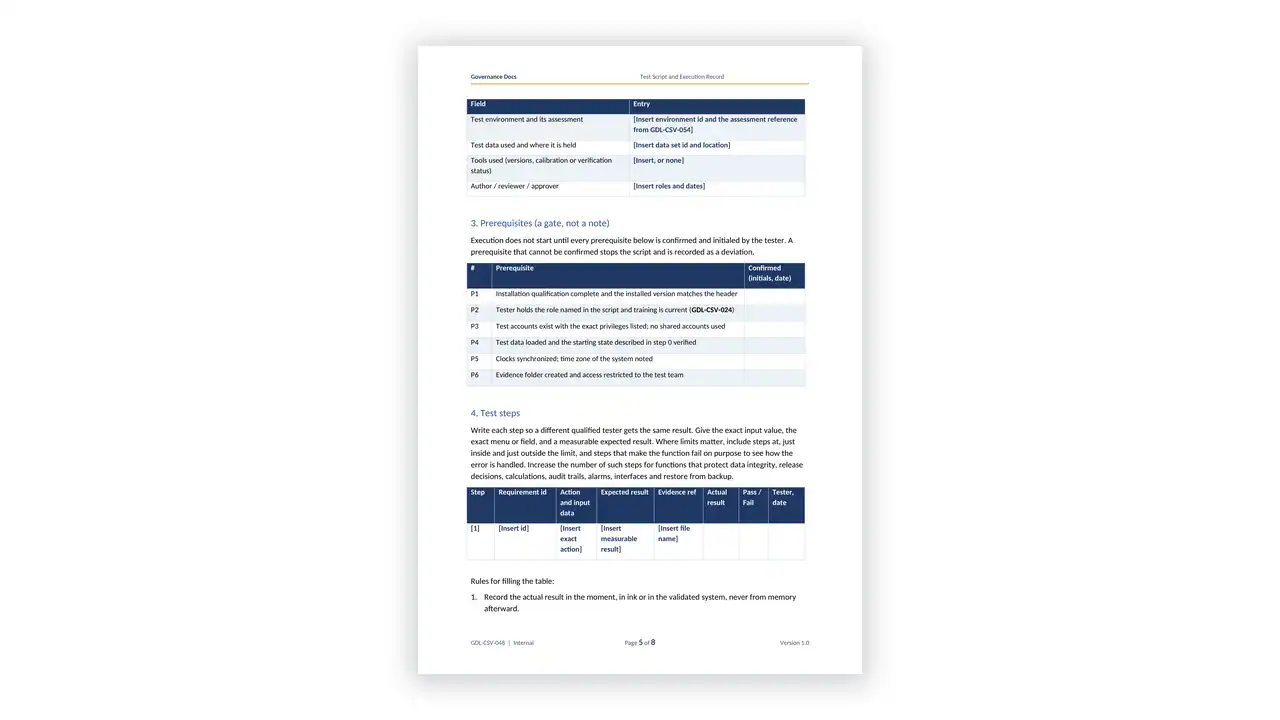
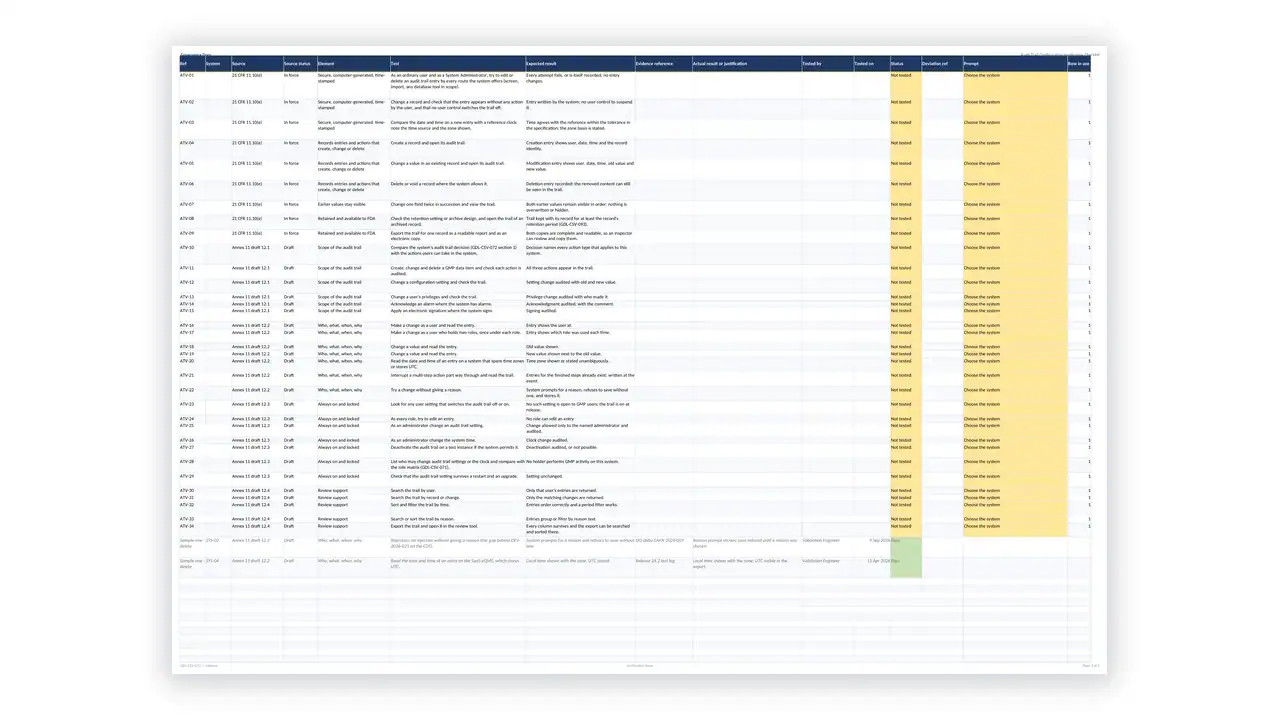
prerequisites, test steps, expected and actual results, tester sign-off and deviation handling
conditional release
System Retirement Plan and Report
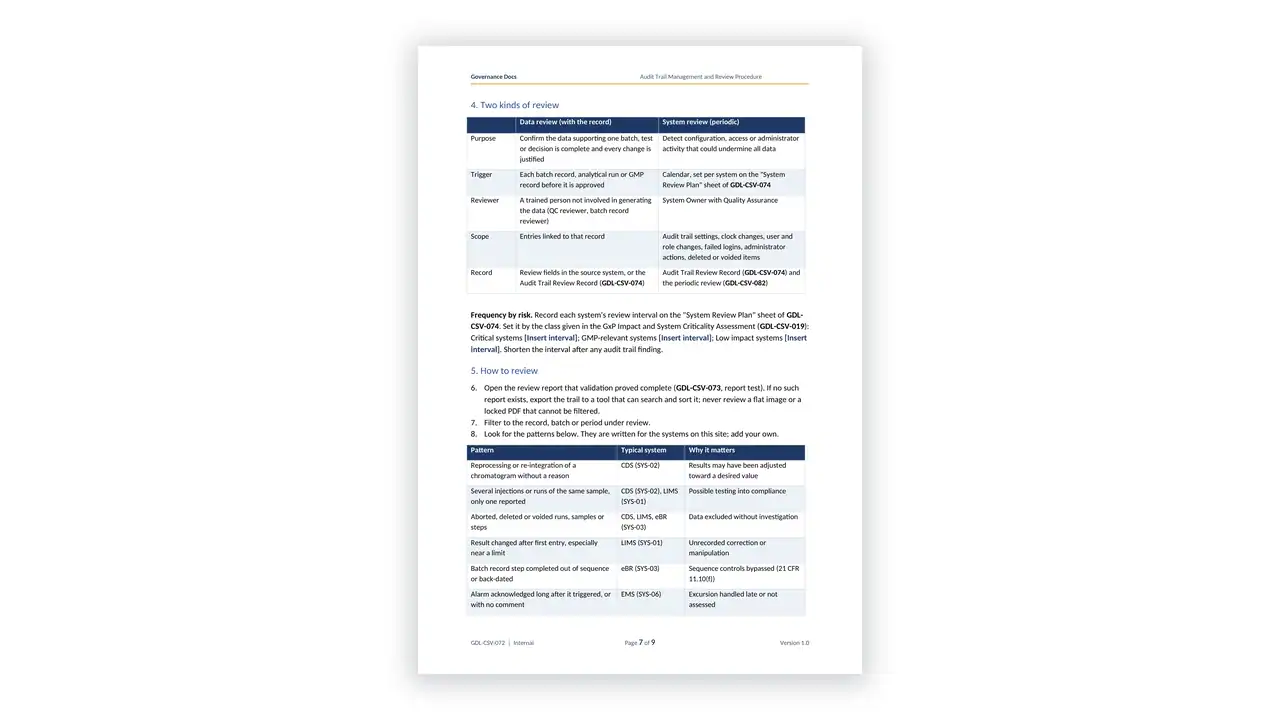
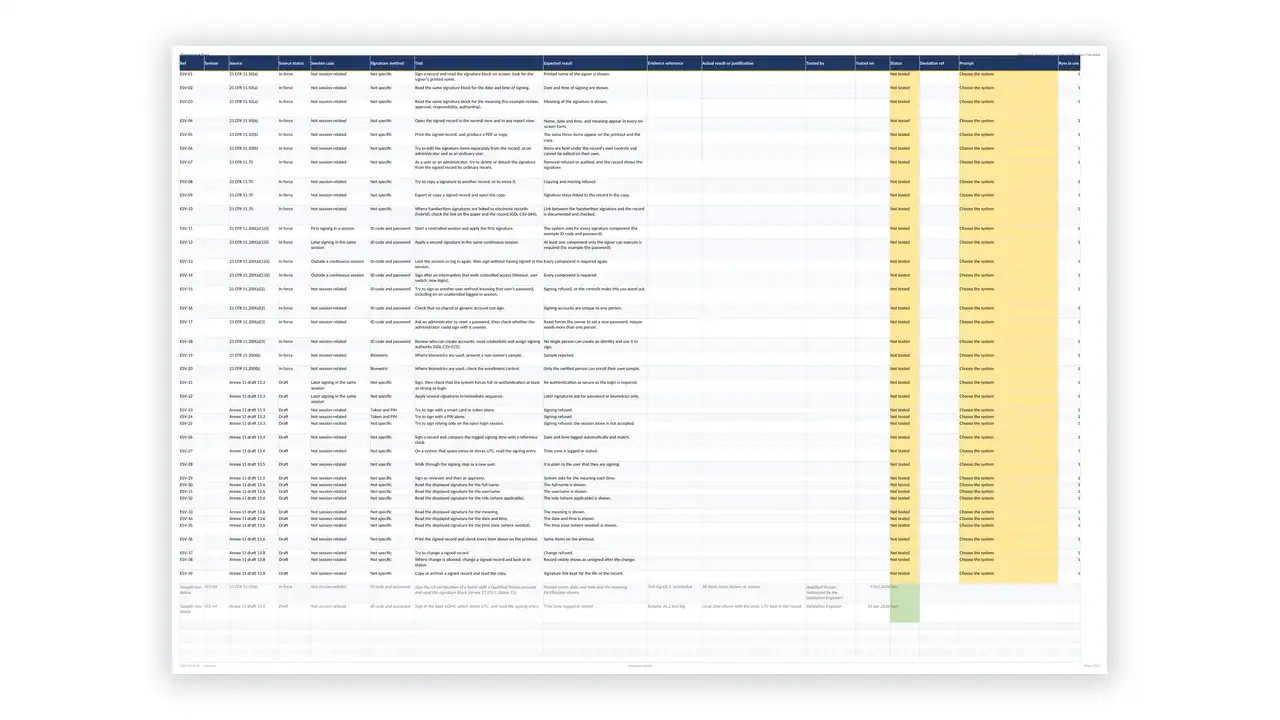
Audit trails get a management and review procedure with a configuration checklist and a review record. Electronic signatures get a procedure, an identity verification record, a controls checklist and the FDA Electronic Signature Certification Letter that 11.100(c) requires before first use. Access, passwords, lost tokens and attempted misuse each have their own document, as do copies and printouts, hybrid records, backup, archiving, periodic review, security and disaster recovery.
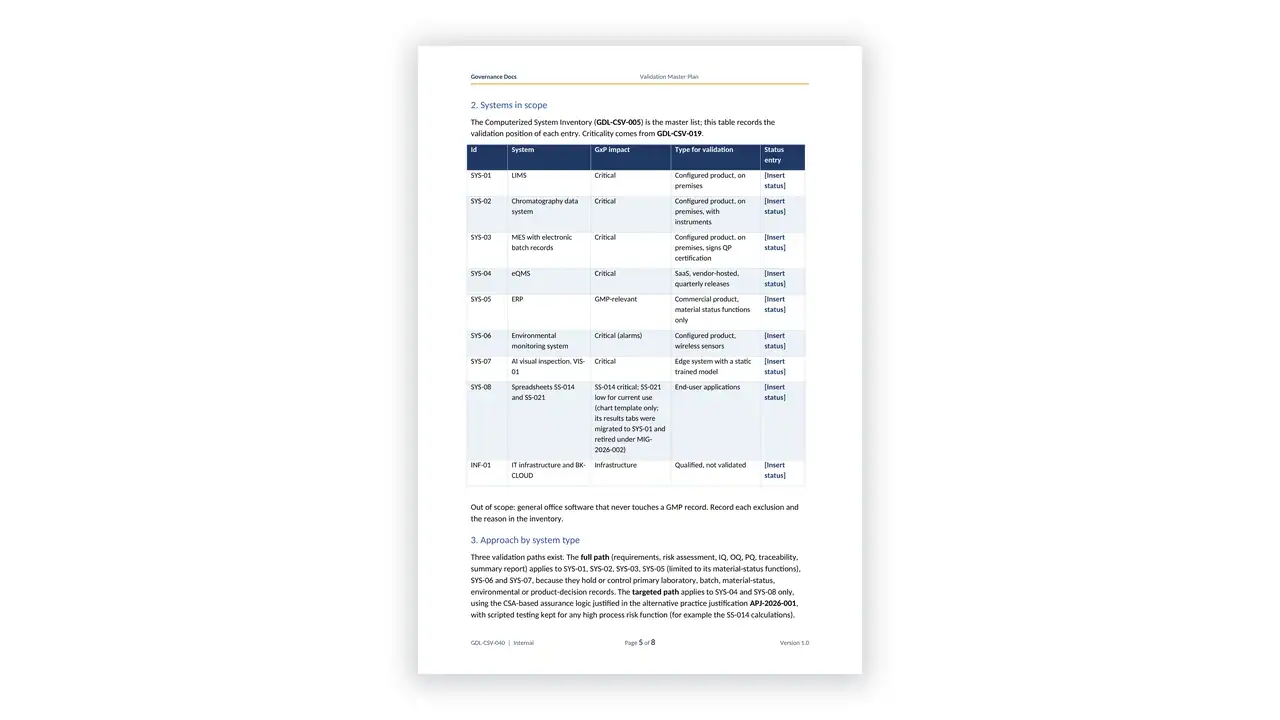
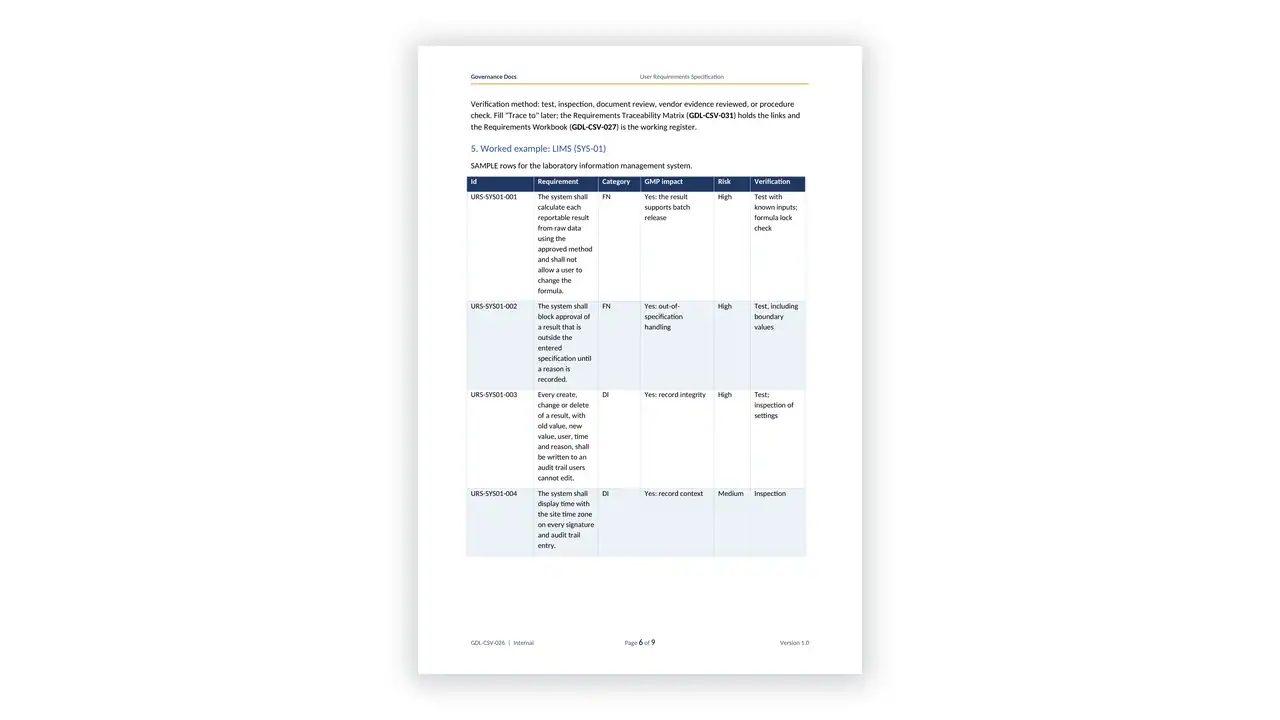
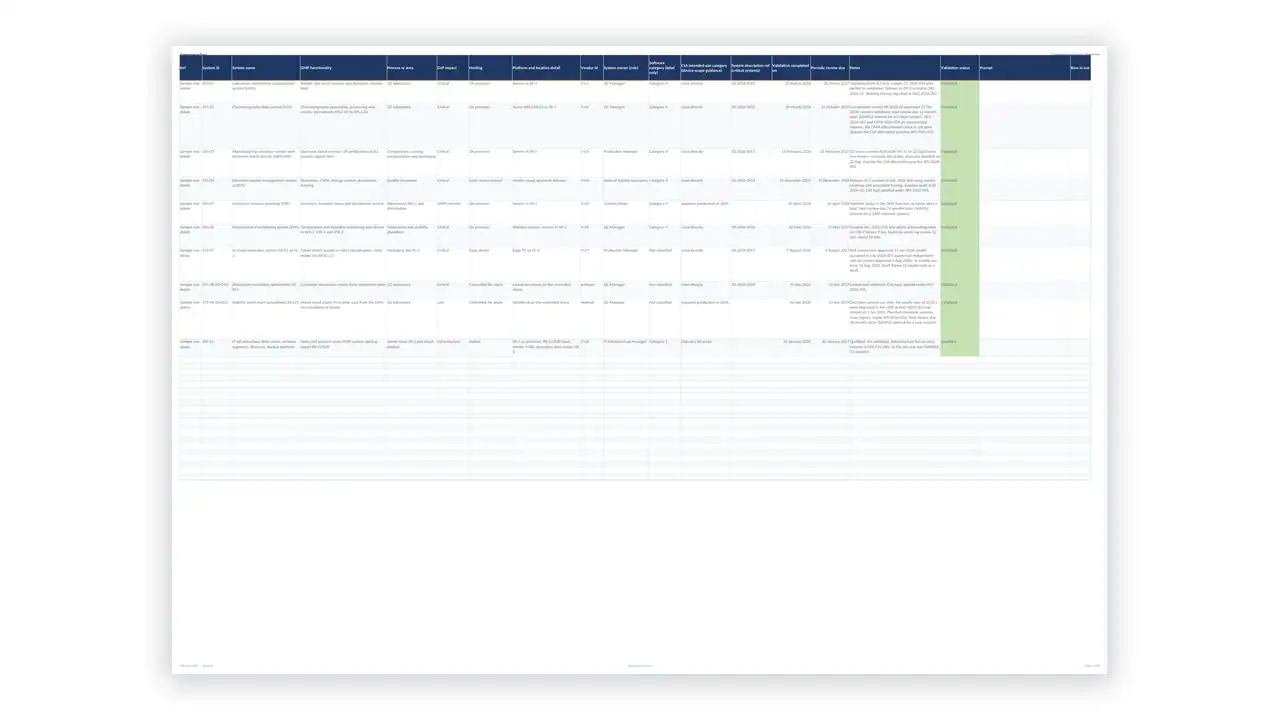
The worked examples follow one fictional tablet and capsule manufacturer that supplies the US and the EU, with a LIMS, a chromatography data system, an MES with electronic batch records, a SaaS quality system, an environmental monitoring system, an AI visual inspection station and controlled spreadsheets. A LIMS upgrade, an audit trail finding and its CAPA, a late alarm acknowledgment, an access review and a disaster recovery test each keep the same reference, dates and facts wherever they appear. Limits your site must set, such as password length or review intervals, stay as placeholders, so a number in an example is never presented as a regulatory requirement.
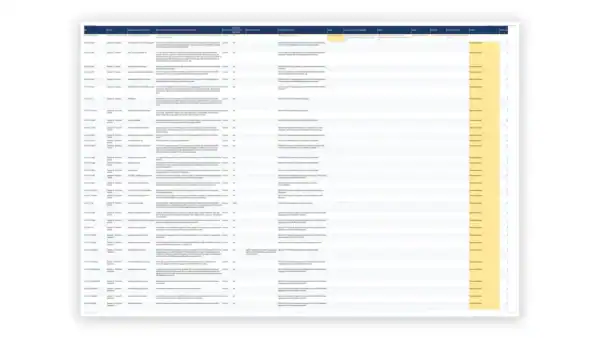
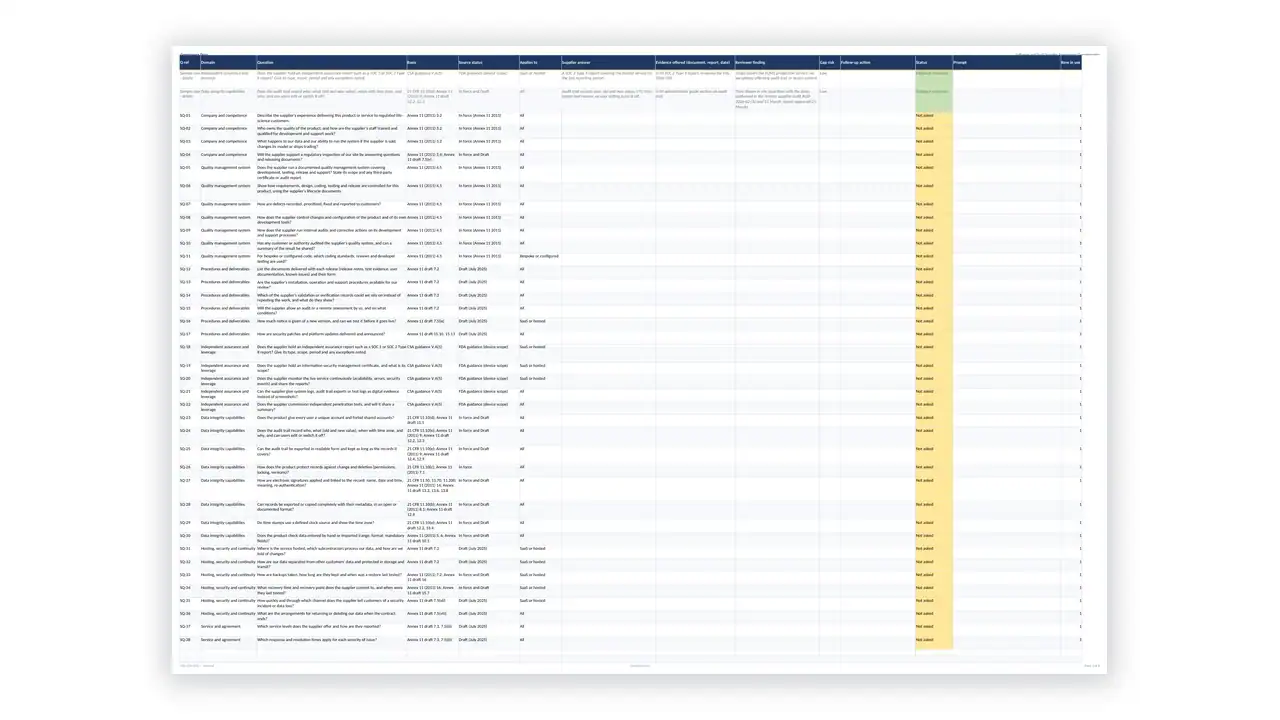
The 37 Excel workbooks calculate what can be calculated: due dates, risk priority numbers and summaries. They never mark a record passed, approved or closed. You choose each row’s Status from a dropdown, and a Prompt column flags anything missing, overdue or inconsistent, for example a reviewer who also created the data, or a GMP user who also holds administrator rights. Rows typed below the ruled range are counted on the summary, so nothing goes missing quietly.
| Folder | Topic | Main references | Documents |
|---|---|---|---|
| 00 | Program Foundation | Part 11 Subpart A; Annex 11 Principle | 6 |
| 01 | Principles and Quality System | Annex 11 draft 2-3; Annex 11 10, 13 | 11 |
| 02 | Risk Management | Annex 11 1; draft 4; CSA V.A(2) | 4 |
| 03 | Personnel and Training | Annex 11 2; draft 5; 11.10(i)(j) | 4 |
| 04 | System Requirements | Annex 11 4.3-4.4; draft 6 | 6 |
| 05 | Supplier and Service Management | Annex 11 3; draft 7 | 6 |
| 06 | Alarms | Annex 11 draft 8 | 2 |
| 07 | Qualification and Validation | 11.10(a); Annex 11 4; draft 9; CSA | 18 |
| 08 | Handling of Data | Annex 11 5, 6, 8; draft 10 | 7 |
| 09 | Identity and Access Management | 11.10(d)(g), 11.300; Annex 11 12; draft 11 | 7 |
| 10 | Audit Trails | 11.10(e); Annex 11 9; draft 12 | 3 |
| 11 | Electronic Signatures | 11.30-11.300; Annex 11 14-15; draft 13 | 6 |
| 12 | Periodic Review | Annex 11 11; draft 14 | 3 |
| 13 | Security and Continuity | Annex 11 12, 16; draft 15 | 7 |
| 14 | Backup and Archiving | 11.10(c); Annex 11 7, 17; draft 16-17 | 4 |
| 15 | AI Models (Draft Annex 22) | Annex 22 draft | 3 |
| 16 | Assessment, Mapping and Inspection | All sources | 8 |
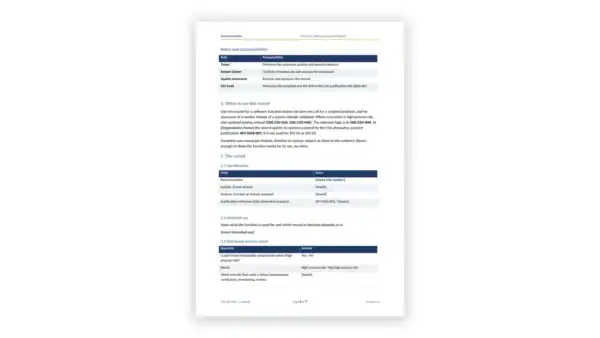
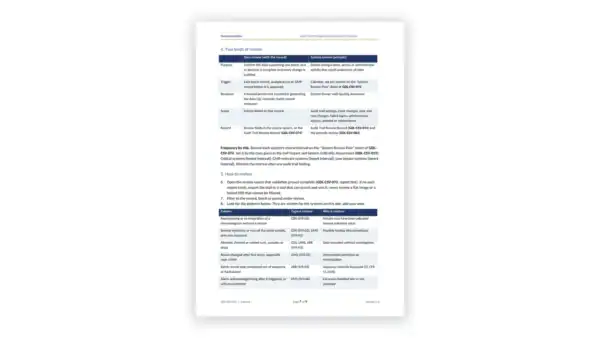
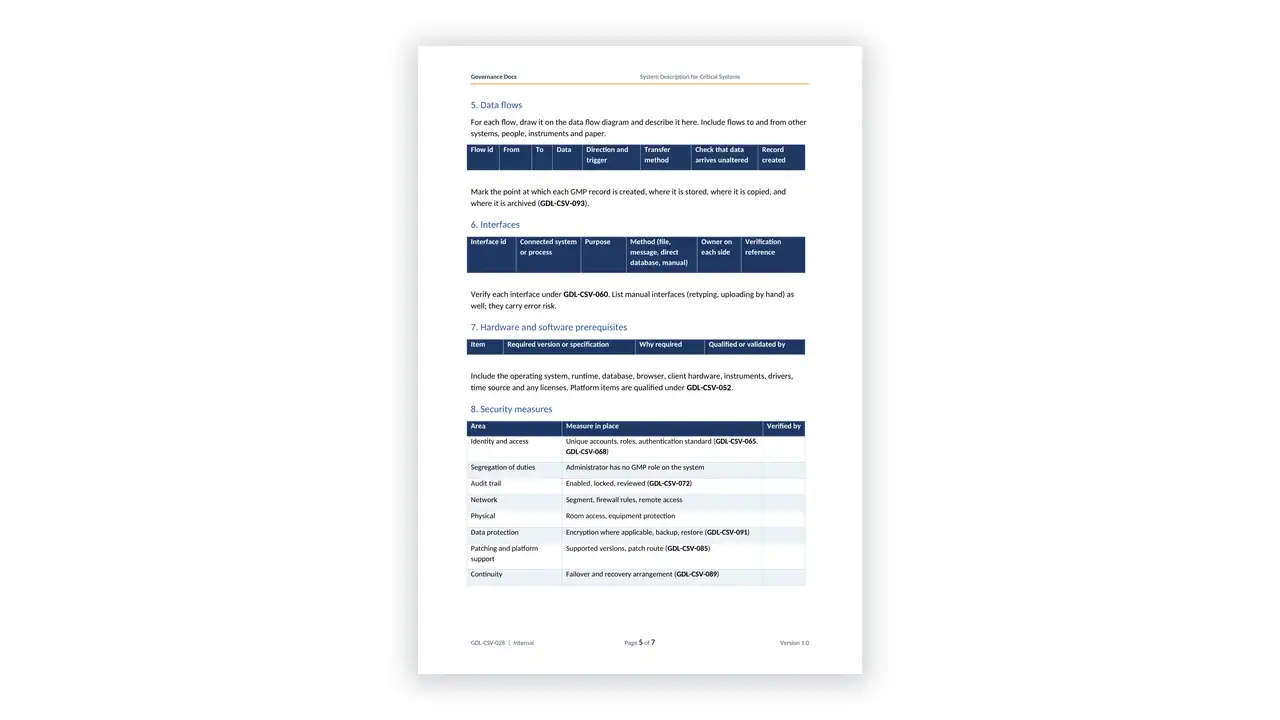
The 21 CFR Part 11 Toolkit is 68 Word documents and 37 Excel workbooks. Every file has a branded cover and the same styling throughout. Every placeholder is a [square-bracket] token, such as [Organization Name], so one find-and-replace makes the templates yours. The documents are written in US English.
21 CFR Part 820, and the ISO 13485 Toolkit the standard it incorporates.
covers the software life cycle; this toolkit covers the software you use to make and control products.
The 21 CFR Part 11 Toolkit is independent of FDA, the European Commission and EMA, and is not approved or endorsed by them. It cites section numbers and short names and describes each requirement in our own words; the shipped files were compared mechanically against the regulatory texts, and no run of words matches beyond headings and references. Read 21 CFR Part 11 on ecfr.gov and Annex 11 in EudraLex Volume 4 alongside the toolkit. GAMP 5 is published by ISPE and is not included; the pack names GAMP software categories as a label only.
00 — Program Foundation (6 documents)
01 — Principles and Quality System (11 documents)
02 — Risk Management (4 documents)
03 — Personnel and Training (4 documents)
04 — System Requirements (6 documents)
05 — Supplier and Service Management (6 documents)
06 — Alarms (2 documents)
07 — Qualification and Validation (18 documents)
08 — Handling of Data (7 documents)
09 — Identity and Access Management (7 documents)
10 — Audit Trails (3 documents)
11 — Electronic Signatures (6 documents)
12 — Periodic Review (3 documents)
13 — Security and Continuity (7 documents)
14 — Backup and Archiving (4 documents)
15 — AI Models (Draft Annex 22) (3 documents)
16 — Assessment, Mapping and Inspection (8 documents)
The 21 CFR Part 11 Toolkit is 105 editable Microsoft Word and Excel templates for computer system validation and electronic records under 21 CFR Part 11 and EU GMP Annex 11: 68 documents and 37 workbooks, organized in 17 folders.
No template can. Compliance depends on how each system is configured, validated and used. The 21 CFR Part 11 Toolkit gives you the procedures, protocols and records to validate your systems and show inspectors the controls; your team still runs them.
Yes, as a draft. Every numbered sub-clause of the July 2025 consultation draft is mapped and labeled as draft, alongside the 2011 text that is in force today. Check EudraLex Volume 4 for the adopted text when it is published.
FDA wrote the CSA guidance for device manufacturers. A drug or biologics site may apply the same risk-based logic only with a documented justification, which the Alternative Practice Justification Record is built for. The validation procedures work with or without it.
No. GAMP 5 is copyright of ISPE. The 21 CFR Part 11 Toolkit refers to GAMP software categories as a label but reproduces none of its text.
Microsoft Word (.docx) and Microsoft Excel (.xlsx), 68 and 37 respectively. Every file is fully editable, unlocked and unprotected. The workbooks have dropdowns, frozen headers and worked example rows that you delete before use.
With the CSV Program Guide and How to Use This Toolkit, then the Part 11 Applicability and Predicate Rule Determination Procedure to decide which of your records and systems are in scope, and the Computerized System Inventory to list them.
Implementing for clients? The Consultant Package bundles 92 toolkits — 8,500+ editable templates — under one firm-wide licence that covers unlimited client engagements. $1,399 one-time.
Free articles that explain the requirements this toolkit covers.
 Comprehensive 21 CFR Part 11 Toolkit – 105 Computer System Validation Templates$99.00








Reviews
There are no reviews yet